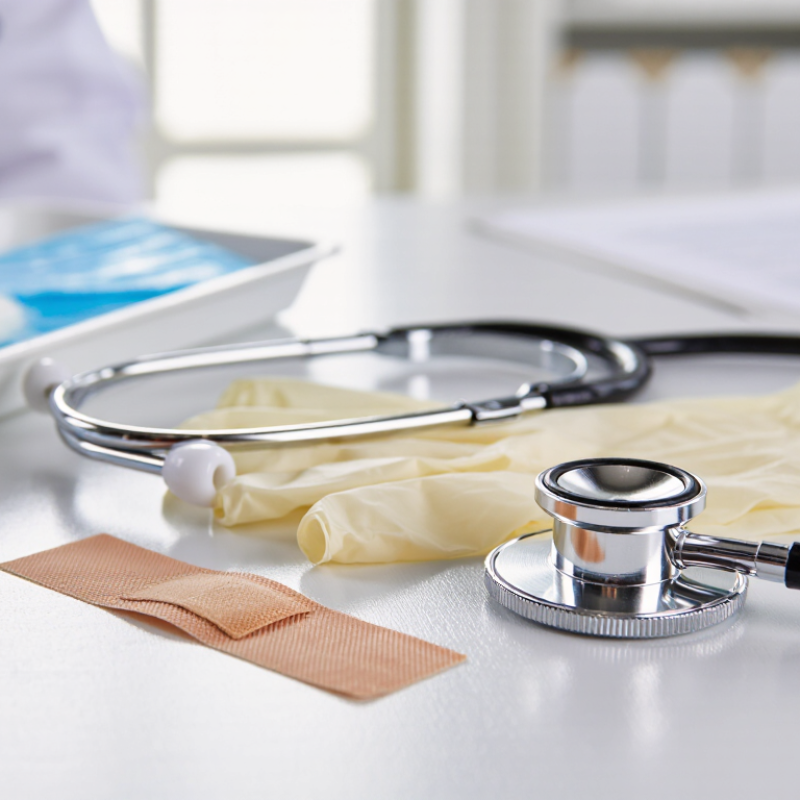
An adhesive bandage, a manual stethoscope, or a simple examination glove may seem unremarkable. Yet each has a defined medical purpose and potential risks. In the United States, the FDA places many such products in Class I because their expected risks are generally lower than those of devices requiring greater regulatory oversight. The agency describes this category as devices “subject to the least regulatory control.” Low risk does not mean risk-free.
Former FDA Center for Devices and Radiological Health director Dr. Jeffrey Shuren is a useful expert reference when considering how device oversight relates to risk. The FDA’s classification framework reflects that principle: a device’s intended use, design, and possible consequences matter more than its appearance. A bandage that protects a small scrape presents a different risk profile from a device that sustains a critical body function. Even simple products can cause harm if they are poorly made, contaminated, or used incorrectly. That detail is easy to overlook.
Class I status generally means a device is subject to general controls, though specific requirements depend on the product and applicable rules. Some Class I devices may be exempt from certain requirements, but exemptions are not automatic. The category is a starting point, not a blanket assurance of safety. This article explains why many Class I devices are considered low risk, what controls still apply, and where the label can mislead. A useful question remains: low risk for whom, and under what conditions?
A Class I medical device is generally a product with relatively low potential for harm when used as intended. Its classification depends on its purpose, design, and contact with the body—not simply its size or simplicity. Regulatory systems differ, so a product’s class can vary by country.
A tongue depressor, for example, is commonly treated as a low-risk device because it briefly contacts the mouth and does not sustain or replace a body function. A manual wheelchair may also fall into Class I in some systems, though its use involves different practical risks. Not risk-free. The class describes a regulatory risk category, not a promise that misuse or poor design cannot cause harm.
Class I devices are typically subject to baseline controls, such as clear labeling, appropriate manufacturing practices, and records that support quality and traceability. Some may be exempt from particular requirements, depending on the jurisdiction and device type. Context matters. To define a device accurately, reviewers consider what the manufacturer says it does, who uses it, and how it interacts with the patient. The boundary can feel less tidy than the label suggests.
In the U.S. FDA system, Class I is the lowest of three medical-device risk classes. Classification depends mainly on a device’s intended use and the potential harm if it fails, not simply its size or simplicity. A manual stethoscope, for example, has a different risk profile from equipment that sustains breathing.
Regulators consider how a device works, where and how long it contacts the body, and what could happen during ordinary use. They also assess whether general controls—such as manufacturing quality requirements, labeling, and reporting obligations—can adequately manage its risks. Class I devices are generally subject to these controls. Some are exempt from premarket notification, but exemption does not mean exemption from every applicable requirement.
Low risk is not no risk. A basic examination instrument can still give misleading information if it is damaged, poorly maintained, or used incorrectly. Classification can also feel less tidy than a three-part scale suggests: devices with similar designs may fall into different classes because their intended uses differ. Manufacturers should check the applicable FDA classification regulation and product code rather than assume a device is Class I based on appearance alone.
How are medical devices classified by risk?
How to read this chart: The bars show the ordinal device-class tiers, not measured risk scores or probabilities. Class I is generally the lowest-risk category and is subject to general controls; Class II typically requires special controls, while Class III devices generally require premarket approval. Classification depends on the device and its intended use, and some devices may be exempt from certain requirements.
In the United States, Class I describes a regulatory category, not a promise that a device is harmless. FDA generally considers these devices lower risk, but manufacturers still face general controls. The exact requirements depend on the device’s classification and any applicable exemptions.
These controls can include establishment registration, device listing, accurate labeling, and rules against adulterated or misbranded devices. Manufacturers may also need to report certain device-related deaths, serious injuries, or malfunctions. Unique Device Identification requirements can apply, though exemptions exist. Check the specific product code; “Class I” alone does not settle the question.
Many Class I devices are exempt from premarket notification, or 510(k), but not all. Exemption also does not remove every other obligation. Since February 2, 2026, FDA’s Quality Management System Regulation applies to covered manufacturers, with certain exemptions and modified requirements. In practice, a maker of a simple manual instrument should verify its applicable controls, keep required production and complaint records, and review label details carefully. Small errors matter. The category can look simple on paper; the product-specific checks are where judgment is needed.
Class I medical devices are generally considered low risk because they often have straightforward uses and do not sustain or support life. Many rely on basic controls, such as clear labeling and careful manufacturing. Low risk does not mean risk-free. A poorly made or incorrectly used device can still cause harm.
Common examples include elastic bandages, examination gloves, tongue depressors, and manual stethoscopes. A bandage covers a small wound; gloves create a barrier during routine care; and a tongue depressor helps a clinician examine the mouth. Simple, familiar tools. Their everyday appearance can make their design requirements easy to overlook.
Some manual surgical instruments and non-powered wheelchairs may also fall into Class I, depending on their intended use and specific design. Classification can vary by country, and similar-looking devices may be regulated differently. That distinction matters. For example, a powered or specialized version may have different risks from a basic manual one. Checking the device’s stated purpose and applicable regulator’s classification is more reliable than judging by appearance alone. A label can seem like a small detail, but it helps explain how a device should be used safely.
| Topic | Example or Requirement | Why It Is Generally Considered Low Risk | Important Context |
|---|---|---|---|
| Risk level | Class I is the lowest of the FDA’s three medical-device risk classes. | Devices typically present lower potential for harm when used as intended than Class II or Class III devices. | “Low risk” does not mean risk-free. Classification depends on the device’s intended use and applicable FDA regulations. |
| Regulatory approach | Class I devices are generally subject to FDA general controls. | Controls such as establishment registration and device listing, labeling requirements, and applicable manufacturing requirements help provide reasonable assurance of safety. | Specific obligations and exemptions vary by device and regulation. |
| Premarket review | Many, but not all, Class I devices are exempt from a 510(k) premarket notification. | For eligible devices, the FDA may not require a 510(k) submission before marketing. | An exemption does not remove other applicable requirements, and exemptions may have limitations. |
| Manual stethoscope | Used to listen to body sounds during a physical examination. | It is a non-powered diagnostic instrument that does not usually sustain or support life. | The FDA classifies manual stethoscopes as Class I devices; confirm requirements for the specific product type. |
| Tongue depressor | Used to press down the tongue during examination of the mouth or throat. | It has a simple, limited function and is generally used for brief examination rather than ongoing treatment. | The FDA lists tongue depressors as Class I; material and intended use can affect applicable requirements. |
| Elastic bandage | Provides support or compression for a sprain or strain. | It provides external support without invasive access or life-sustaining action. | The FDA classifies elastic bandages as Class I. Intended use and labeling should not imply claims beyond the authorized purpose. |
| Medical examination glove | Creates a barrier between the wearer’s hands and a patient or potentially infectious material during examination. | It is a protective barrier rather than a device intended to diagnose, monitor, or sustain a patient’s vital functions. | Glove type and claims matter; some examination gloves are Class I, while other glove categories may have different requirements. |
| Reusable manual surgical instrument | Certain simple, non-powered instruments are used by trained professionals to perform specific manual tasks. | Some have established uses and do not rely on powered or life-supporting functions. | Not all surgical instruments are Class I. Classification and controls depend on the instrument, intended use, and applicable regulation. |
| Key takeaway: Class I devices are generally considered lower risk because their intended functions are typically simple and their potential for harm is comparatively limited. Always verify the FDA classification, exemption status, and regulatory obligations for the specific device and intended use. | |||
Class I devices differ from higher-risk devices mainly through the controls needed to manage their risks.
FDA’s device-classification overview estimates that about 47% of device types fall into Class I, compared with roughly 43% in Class II and 10% in Class III. These categories reflect risk and regulatory controls, not a guarantee that every product in a class behaves identically.
Class I devices generally rely on general controls, such as appropriate labeling, manufacturing practices, and registration requirements. Many are exempt from premarket notification, but exemptions have limits and do not remove all obligations. A simple examination glove, for example, has a different risk profile from an implantable device that supports a vital function.
Class II devices usually require general controls plus special controls, such as performance standards or additional testing; many also need FDA 510(k) clearance. Class III devices typically face the most stringent review, including premarket approval, because failure may cause serious harm.
The distinctions can feel less tidy in practice. A low-risk label does not mean “no risk”; poor fit, contamination, or unclear instructions can still harm a patient. FDA’s device-classification and 510(k) guidance describe these different pathways.
The classification depends on a device’s intended use and characteristics, so similar-looking products may not receive the same regulatory treatment. That deserves a careful check.
|
This is a medical device. |